Cystic Fibrosis
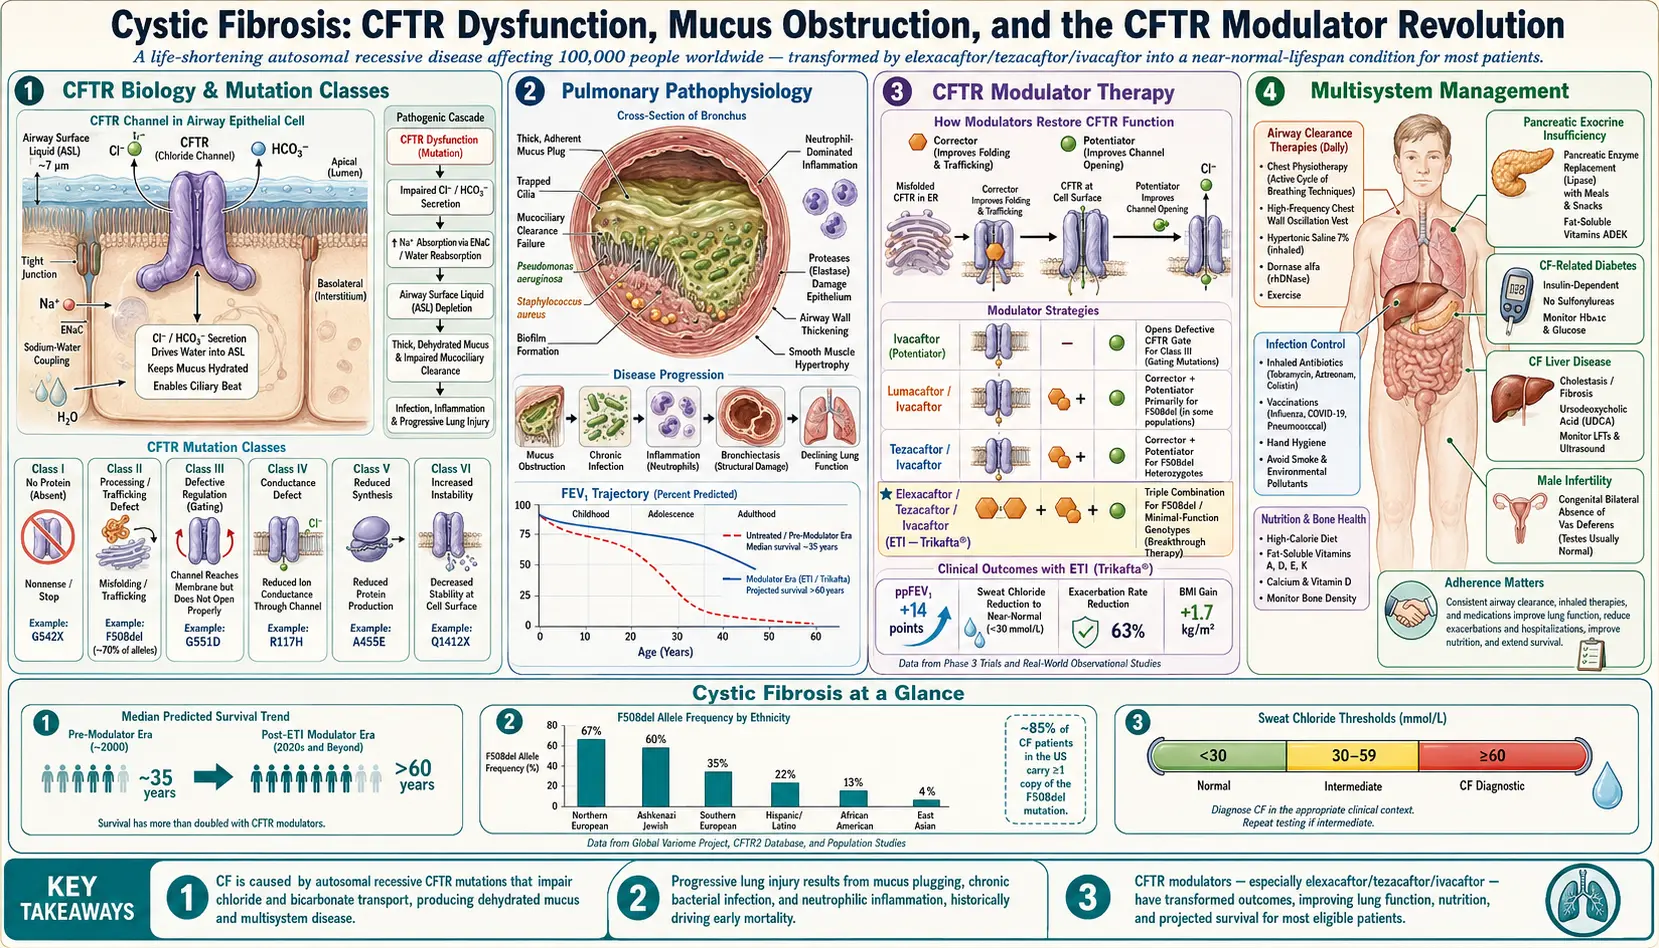
Table of Contents
What Cystic Fibrosis Is
Cystic fibrosis (CF) is a genetic disorder that primarily affects the lungs and digestive system. It is caused by mutations in the CFTR (cystic fibrosis transmembrane conductance regulator) gene, leading to the production of thick, sticky mucus that can clog the airways and trap bacteria, resulting in repeated lung infections and breathing difficulties. CF can also affect the pancreas, liver, and other organs, leading to a variety of health complications.
Causes of Cystic Fibrosis
- Genetic inheritance: CF is an autosomal recessive disorder, which means a person must inherit two defective CFTR genes (one from each parent) to develop the disease.
- CFTR gene mutation: This mutation disrupts the function of the CFTR protein, affecting the transport of salt and water in and out of cells, resulting in thick mucus formation.
Common Symptoms of Cystic Fibrosis
- Persistent coughing with thick mucus.
- Frequent lung infections, such as pneumonia or bronchitis.
- Wheezing or shortness of breath
- Salty-tasting skin
- Poor growth and weight gain despite adequate food intake.
- Digestive issues: Including greasy, bulky stools and constipation.
- Sinus infections and nasal polyps.
Complications of Cystic Fibrosis
- Chronic lung infections: Due to mucus buildup that traps bacteria.
- Bronchiectasis: Damage to the airways that makes it difficult to clear mucus.
- Pancreatic damage: Thick mucus can block ducts in the pancreas, affecting enzyme release and nutrient absorption.
- Diabetes: CF-related diabetes can develop due to damage to the pancreas.
- Liver disease: Blocked bile ducts can lead to liver problems.
- Infertility in males: Due to the absence or blockage of the vas deferens.
Diagnosis
- Newborn screening: CF is often diagnosed shortly after birth through blood tests.
- Sweat test: Measures the amount of salt in sweat; high salt levels indicate CF.
- Genetic testing: Identifies mutations in the CFTR gene.
- Chest X-rays and computed tomography (CT) scans: To assess lung damage and detect complications.
- Lung function tests: Evaluate how well the lungs are working.
Treatment Options
- Airway clearance techniques: Includes chest physiotherapy and devices that help remove mucus from the lungs.
- Medications:
- Bronchodilators: Open the airways to improve breathing.
- Mucus-thinning drugs: Help thin mucus so it can be coughed out more easily.
- Antibiotics: Used to treat and prevent lung infections.
- CFTR modulators: Drugs that improve the function of the defective CFTR protein — ivacaftor (a “potentiator”); lumacaftor-ivacaftor and tezacaftor-ivacaftor, in which a “corrector” is given together with ivacaftor; and the newer triple combinations elexacaftor-tezacaftor-ivacaftor and vanzacaftor-tezacaftor-deutivacaftor.
- Enzyme supplements: Aid in digestion and nutrient absorption.
- High-calorie diet: Necessary to maintain energy levels and support growth.
- Lung transplant: Considered for severe cases when lung function is significantly compromised.
Triple-combination trials. Nearly 90% of people with cystic fibrosis carry at least one copy of the most common CFTR mutation, Phe508del (also written F508del). In a 2019 trial of 403 people aged 12 and older with one Phe508del copy and a second, minimal-function mutation, elexacaftor-tezacaftor-ivacaftor gave lung function (FEV1, the air blown out in one second, as a percentage of the expected value) 14.3 percentage points higher than placebo through 24 weeks, 63% fewer flare-ups (pulmonary exacerbations), and sweat chloride 41.8 mmol per liter lower; most side effects were mild or moderate, and 1% stopped the drug because of them.
Vanzacaftor-tezacaftor-deutivacaftor is taken once a day, against twice a day for elexacaftor-tezacaftor-ivacaftor. In two 52-week phase 3 trials published in 2025 (971 people aged 12 and older), it matched elexacaftor-tezacaftor-ivacaftor on lung function (a difference of 0.2 percentage points), with mostly mild or moderate side effects. A 2026 open-label study in 67 children aged 2–5 found that 92% reached a sweat chloride below the cystic fibrosis diagnostic threshold of 60 mmol/L after 24 weeks; no child stopped because of side effects. The triple-combination trials described here were all funded by the drugs’ maker.
Prevention and Management Strategies
- Regular follow-ups: Frequent check-ups to monitor and manage symptoms and complications.
- Good nutrition: High-calorie, nutrient-dense foods to support growth and energy needs.
- Exercise: Regular physical activity to improve lung function.
- Hydration: Helps keep mucus thin and easier to expel.
- Smoke and pollutants: Exposure raises the risk of lung infections and irritation.
Living with Cystic Fibrosis
Advancements in treatment have significantly improved the quality of life and life expectancy for people with cystic fibrosis. Early diagnosis, comprehensive treatment plans, and supportive care can help manage symptoms, reduce complications, and enable individuals to live fulfilling lives.
A 2023 computer-model study projected a median survival of 71.6 years for people with two copies of F508del who take elexacaftor-tezacaftor-ivacaftor from age 12 or older — 33.5 years longer than with supportive care alone — and 82.5 years when the drug is started between ages 12 and 17. These figures are projections from a model, not observed survival.
Key Research Papers
- Elborn JS. Cystic fibrosis. Lancet. 2016;388(10059):2519-2531. — doi:10.1016/s0140-6736(16)00576-6
- Riordan JR, Rommens JM, Kerem B, et al. Identification of the cystic fibrosis gene: cloning and characterization of complementary DNA. Science. 1989;245(4922):1066-1073. — doi:10.1126/science.2475911
- Middleton PG, Mall MA, Dřevínek P, et al. Elexacaftor-Tezacaftor-Ivacaftor for Cystic Fibrosis with a Single Phe508del Allele. N Engl J Med. 2019;381(19):1809-1819. — doi:10.1056/NEJMoa1908639
- Ramsey BW, Davies J, McElvaney NG, et al. A CFTR potentiator in patients with cystic fibrosis and the G551D mutation (STRIVE). N Engl J Med. 2011;365(18):1663-1672. — doi:10.1056/NEJMoa1105185
- Wainwright CE, Elborn JS, Ramsey BW, et al. Lumacaftor-Ivacaftor in Patients with Cystic Fibrosis Homozygous for Phe508del CFTR (TRAFFIC and TRANSPORT). N Engl J Med. 2015;373(3):220-231. — PubMed PMID: 25981758
- Mogayzel PJ Jr, Naureckas ET, Robinson KA, et al. Cystic fibrosis pulmonary guidelines: chronic medications for maintenance of lung health. Am J Respir Crit Care Med. 2013;187(7):680-689. — doi:10.1164/rccm.201207-1160oe
- Fuchs HJ, Borowitz DS, Christiansen DH, et al. Effect of aerosolized recombinant human DNase on exacerbations of respiratory symptoms and on pulmonary function in patients with cystic fibrosis. N Engl J Med. 1994;331(10):637-642. — doi:10.1056/nejm199409083311003
- Saiman L, Marshall BC, Mayer-Hamblett N, et al. Azithromycin in patients with cystic fibrosis chronically infected with Pseudomonas aeruginosa. JAMA. 2003;290(13):1749-1756. — doi:10.1001/jama.290.13.1749
- Heijerman HGM, McKone EF, Downey DG, et al. Efficacy and safety of the elexacaftor plus tezacaftor plus ivacaftor combination regimen in people with cystic fibrosis homozygous for the F508del mutation. Lancet. 2019;394(10212):1940-1948. — doi:10.1016/S0140-6736(19)32597-8
- Taylor-Cousar JL, Munck A, McKone EF, et al. Tezacaftor-ivacaftor in patients with cystic fibrosis homozygous for Phe508del (EVOLVE). N Engl J Med. 2017;377(21):2013-2023. — doi:10.1056/nejmoa1709846
- Cystic Fibrosis Foundation Patient Registry. 2020 Annual Data Report. Bethesda, MD: Cystic Fibrosis Foundation. 2021. Find on PubMed
- Cutting GR. Cystic fibrosis genetics: from molecular understanding to clinical application. Nat Rev Genet. 2015;16(1):45-56. — doi:10.1038/nrg3849
- Keating C, Yonker LM, Vermeulen F, et al. Vanzacaftor-tezacaftor-deutivacaftor versus elexacaftor-tezacaftor-ivacaftor in individuals with cystic fibrosis aged 12 years and older (SKYLINE Trials VX20-121-102 and VX20-121-103): results from two randomised, active-controlled, phase 3 trials. Lancet Respir Med. 2025;13(3):256-271. — PubMed PMID: 39756424
- Rayment JH, Davies G, Gaffin JM, et al. Safety and efficacy of vanzacaftor-tezacaftor-deutivacaftor in children with cystic fibrosis aged 2-5 years (TIMBERLINE Trial VX21-121-105): a phase 3, open-label study. Lancet Respir Med. 2026 (online ahead of print). — PubMed PMID: 42710511
- Lopez A, Daly C, Vega-Hernandez G, MacGregor G, Rubin JL. Elexacaftor/tezacaftor/ivacaftor projected survival and long-term health outcomes in people with cystic fibrosis homozygous for F508del. J Cyst Fibros. 2023;22(4):607-614. — PubMed PMID: 36849331
Historical Background
Cystic fibrosis was first recognized as a distinct disease by Dorothy Hansine Andersen in 1938, who described its pathology in a landmark paper connecting pancreatic lesions with lung disease in children. The underlying genetic defect in the CFTR gene was identified in 1989 by Lap-Chee Tsui, John Riordan, and Francis Collins, opening the door to targeted molecular therapies.
PubMed Topic Searches
The following PubMed topic searches surface the current peer-reviewed literature on Cystic Fibrosis. Each link opens a live PubMed query; results update as new papers are indexed.
- PubMed: Search: cystic fibrosis CFTR
- PubMed: Search: cystic fibrosis modulator therapy
- PubMed: Search: ivacaftor tezacaftor elexacaftor
- PubMed: Search: cystic fibrosis lung transplantation
- PubMed: Search: cystic fibrosis pancreatic insufficiency
- PubMed: Search: cystic fibrosis sweat chloride test
- PubMed: Search: cystic fibrosis newborn screening
- PubMed: Search: cystic fibrosis pseudomonas aeruginosa
- PubMed: Search: cystic fibrosis nutritional management
- PubMed: Search: cystic fibrosis related diabetes
- PubMed: Search: cystic fibrosis airway clearance
- PubMed: Search: cystic fibrosis gene therapy
Connections
- Pulmonology
- CRISPR: How Gene Editing Works — interactive animation
- Inheritance: Dominant, Recessive and Punnett Squares — interactive animation
- Your Airway’s Self-Cleaning Escalator (& the Cough) — interactive animation
- Pneumonia
- Diabetes
- Chronic Obstructive Pulmonary Disease
- Vitamin D3
- Vitamin E
- Vitamin A
- Vitamin K
- Pseudomonas Aeruginosa
- Liver Disease
- Shortness of Breath
- Pancreatitis
- Infertility
- Interstitial Lung Disease
- Pulmonary Hypertension
- Inflammatory Markers
- Cirrhosis
- Constipation
- Chronic Cough
- Günter Blobel — why the F508del protein is destroyed before it arrives — and why its drugs are correctors
- Zinc